IMPLANT FILES
Medical device makers have gotten worse at disclosing when their products harm people, new study finds. The FDA let them.
The agency allowed some companies to send in “incomplete, inaccurate, and delayed” information, resulting in an undercount of at least 12,000 adverse events, researchers said.

A new study reveals how data transparency for medical device malfunctions has eroded in recent years, with dangerous consequences for patients. The study, published in the peer-reviewed medical journal The BMJ, found that device makers have increasingly relied on “summary reporting” of medical device practice harms in federally required disclosures — a practice the U.S. Food and Drug Administration said it ended years ago. But the FDA has quietly continued summary reporting through exemptions that were never made public.
The study found that the summary reports are rife with “incomplete, inaccurate, and delayed” information. Fifty-seven percent of reports described an adverse event “without identified device or use problem,” 38 percent selected “insufficient information” in the data field for the device problem, and 11 percent incorrectly described the report as containing only one adverse event, causing an undercount of nearly 12,000 adverse events.
The Implant Files, a 2019 investigation led by the International Consortium of Investigative Journalists, exposed the harms done by poorly-tested medical devices, which often enter the market with little regulation. The project brought together more than 250 journalists in 36 countries. ICIJ found that over 1.7 million injuries and nearly 83,000 deaths were suspected of being linked to medical devices in the U.S. from 2008 to 2017.
One way these harms continued to spread, the investigation found, was bad data. For decades, the FDA allowed device makers to file incident reports through a program known as “alternative summary reporting,” which allowed them to submit quarterly spreadsheets rather than report incidents individually. In practice, this program concealed severe injuries — preventing the public from understanding the risks associated with the devices. The FDA ended the program in 2019, saying in a press release that it was doing so “in the spirit of public transparency.”
The study found that summary reporting for medical devices has continued under exemptions issued by the FDA without any regulation or public notice. The number of summary reports filed under these exemptions more than doubled from 2020 to 2024, from 808 to 1,757 reports. The study analyzed 6,303 adverse event reports filed from July 2019 to June 2025 using summary reporting exemptions. The reports contained a total of 159,503 individual adverse events — almost all serious injury or death.
Madris Kinard, a former FDA official and a co-author of the study, first became aware of the existence of these exemptions in 2019, and has been filling Freedom of Information Act requests for years on the topic.
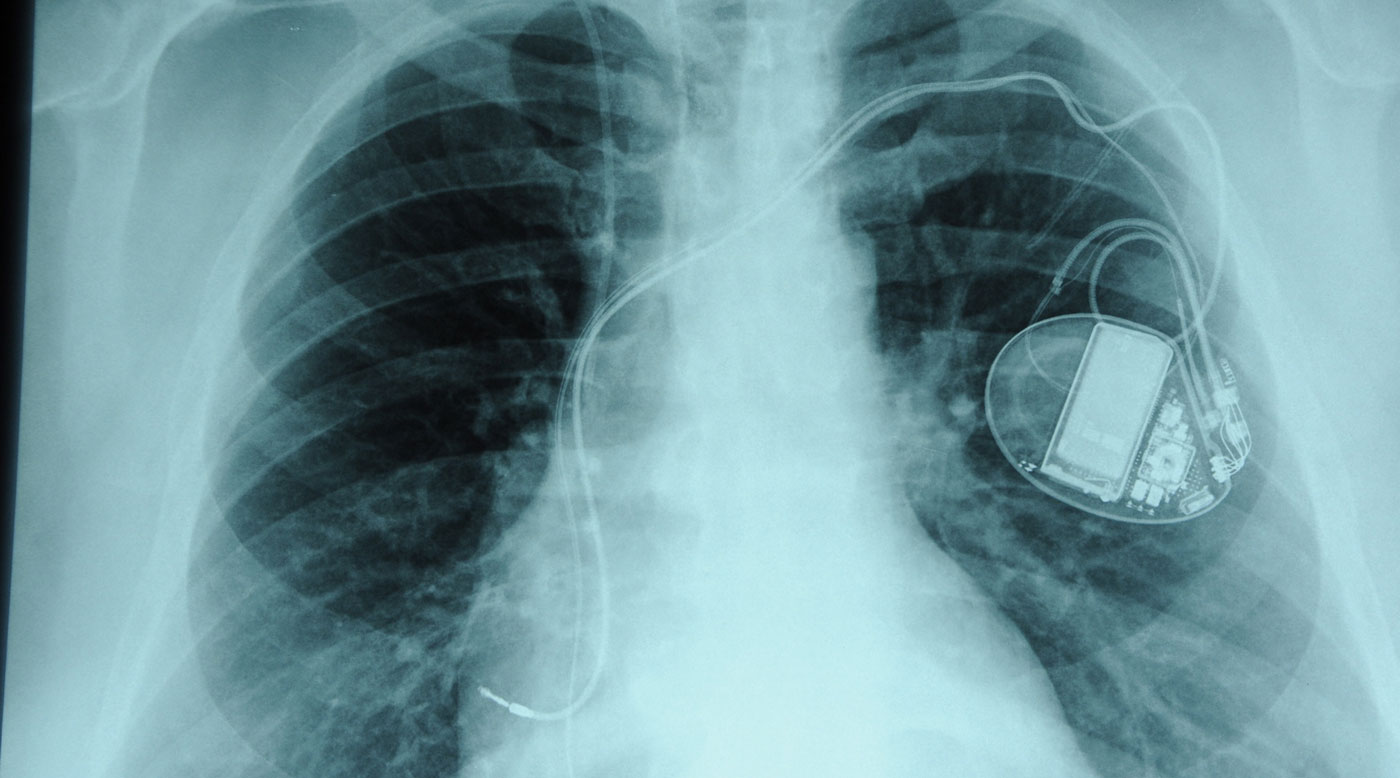
On a call with ICIJ, Kinard pointed to one summary report that catalogued 4,445 deaths of patients using a heart implant designed to reduce the risk of strokes. The report states that “multiple event terms may apply to a single patient.” “You can’t die more than once,” Kinard noted.
Almost all of the adverse events included in the summary reports related to cardiac devices, and over 90 percent of the events were associated with “Class 3” devices, which constitute the highest-risk category regulated by the FDA.
Who is responsible for connecting the dots when a safety signal emerges?
— Madris Kinard, study co-author and former FDA official
The lack of good data on device malfunctions has real-world consequences for patient safety. Nine of the 13 devices that have received summary reporting exemptions have experienced recalls, the study found. These recalls “may have been delayed because of the difficulties of identifying individual adverse event counts and narratives from summary reports,” the study stated.
This issue of data transparency around medical devices burst into public view last week, when the Boston Globe published an investigation on a heart implant that generates nearly $2 billion in annual sales for its manufacturer, Boston Scientific. The company invested tens of millions of dollars in television advertising to consumers, and even paid doctors $30 million to promote the device and train other physicians. But doctors told The Boston Globe that the implant is not more effective than blood thinners, and warned that it was being used in a broader cohort of patients than it was originally intended.
“If physicians cannot see broader safety patterns, researchers cannot independently investigate them, and manufacturers may have different access than regulators, who is responsible for connecting the dots when a safety signal emerges?” Kinard told ICIJ. “Is restricting access to timely, de-identified, device-specific safety information really in the public interest?”